Lesson Overview
This video, presented by Tina Amini (Director of Medtech Combinations), explains the regulatory pathway for combination products under the EU MDR.
Regulatory Classification of Combination Products
- The EU does not have a specific “combination product” classification.
- Instead, products are regulated as either medicinal products or medical devices.
- Primary Mode of Action (PMOA) determines classification:
- Medical Device: Works through physical, simple chemical, or digital means.
- Medicinal Product: Works through pharmacological, metabolic, or immunological action.
Single Integral Drug-Device Combinations Regulated as Medical Devices
- These are devices that incorporate an ancillary medicinal substance.
- MDR Annex VIII, Rule 14 applies:
- Class III (highest risk classification) for devices with medicinal components.
- The rule omits the phrase “liability to act” meaning all such devices fall under Class III.
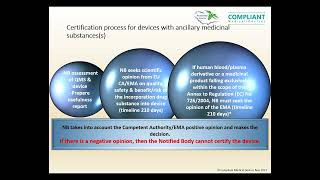
Certification Process for Medical Devices with Medicinal Substances
- The manufacturer applies to a Notified Body, which assesses:
- Quality Management System (QMS) and technical documentation.
- Usefulness of the medicinal substance in the device.
- The Notified Body must consult either:
- EU Competent Authorities or EMA (European Medicines Agency).
- EMA consultation is required if the substance comes from human blood/plasma or falls under Regulation 726/2004.
- Timelines: The consultation process takes at least 210 days, excluding pauses for additional queries.
- Final Decision:
- If the opinion is positive, the Notified Body issues certification.
- If the opinion is negative, certification cannot be granted.
Impact of Article 117 on Drug-Device Combinations Regulated as Medicinal Products
- Article 117 of MDR amends the Medicinal Product Directive.
- Marketing Authorization Applications (MAAs)must include:
- Results of conformity assessment OR
- Notified Body certification (CE marking) OR
- Notified Body opinion report if CE marking is unavailable.
- If the device part is not CE marked, pharmaceutical/biotech companies must:
- Engage with a Notified Body to obtain a Notified Body Opinion (NBOp) on the device component.
- Include the NBOp in the marketing authorisation dossier submitted to the EMA.